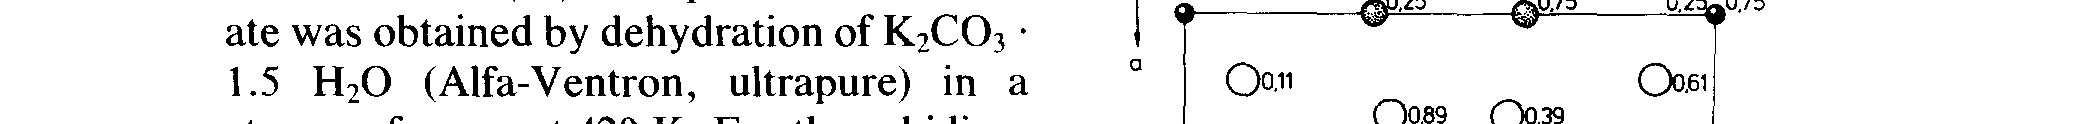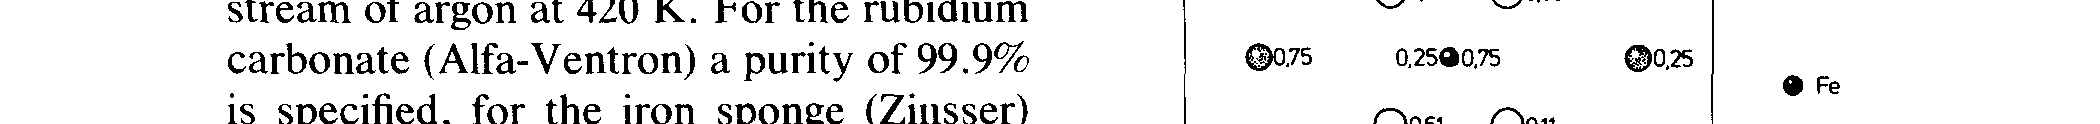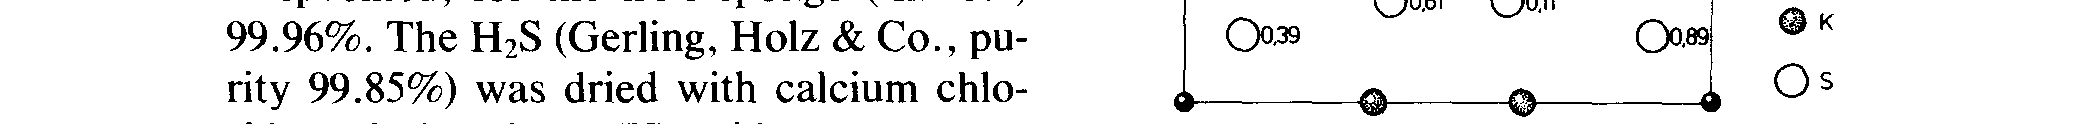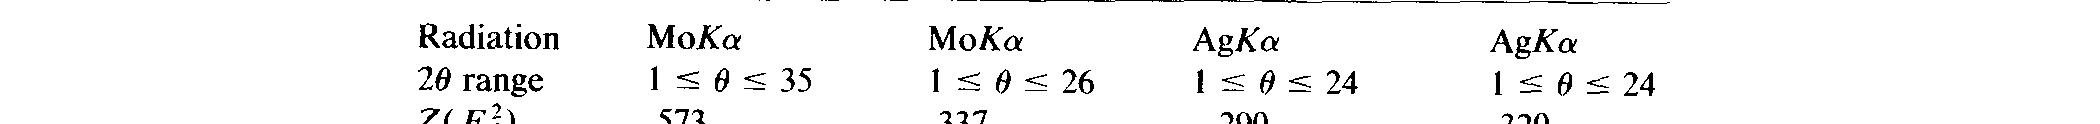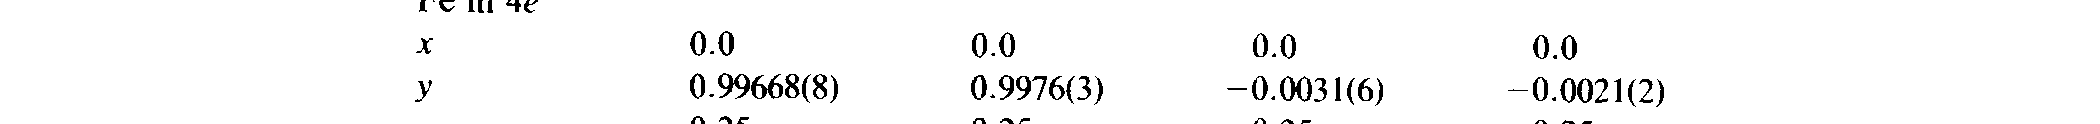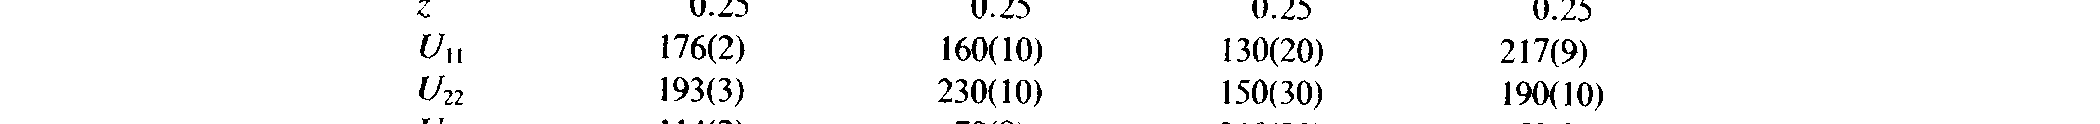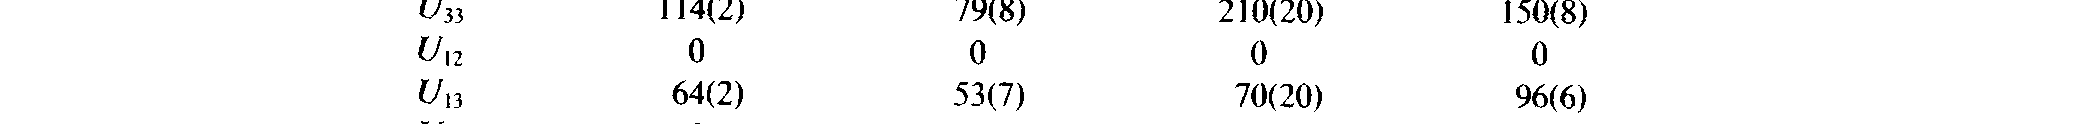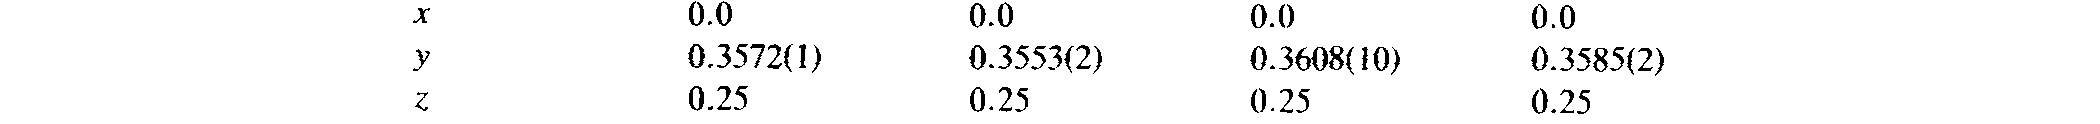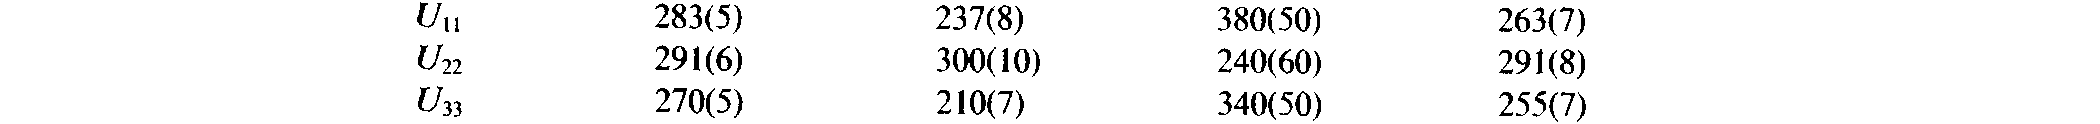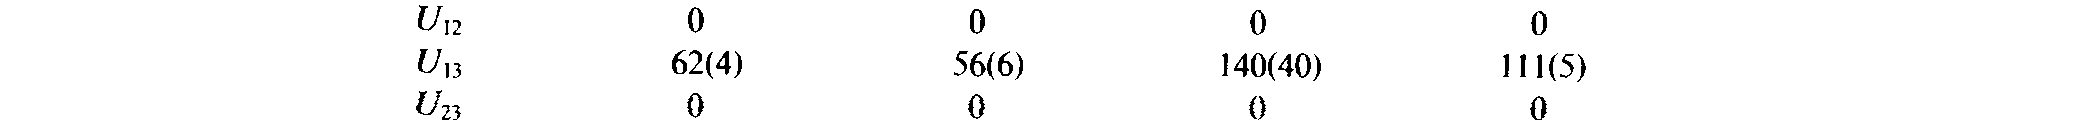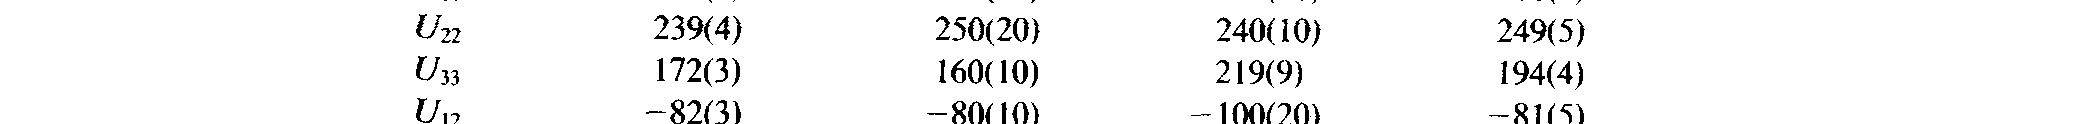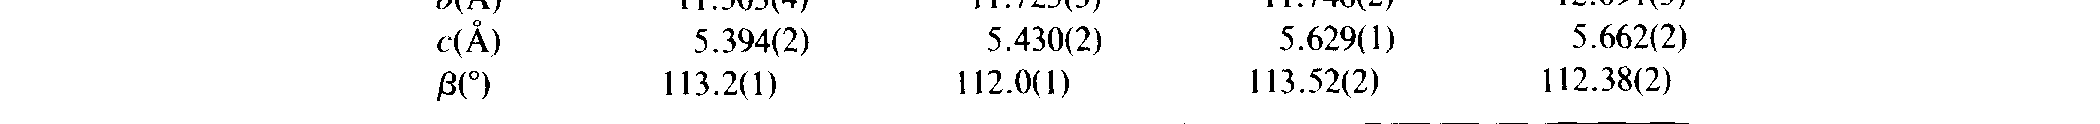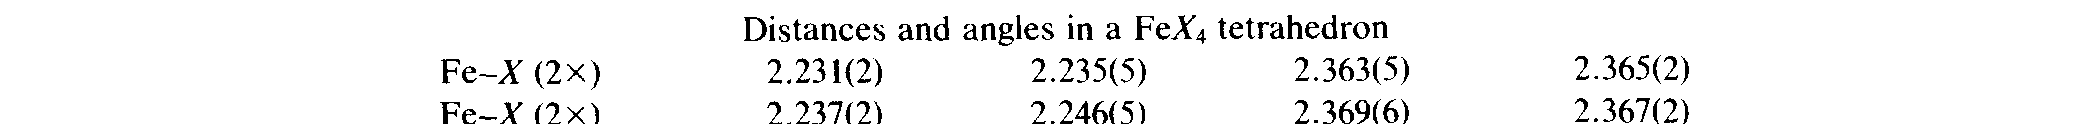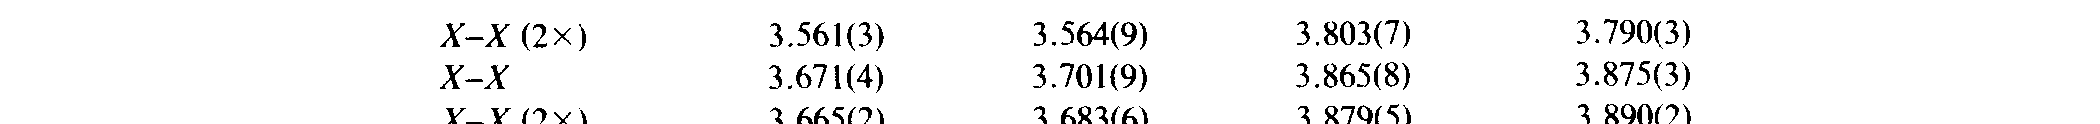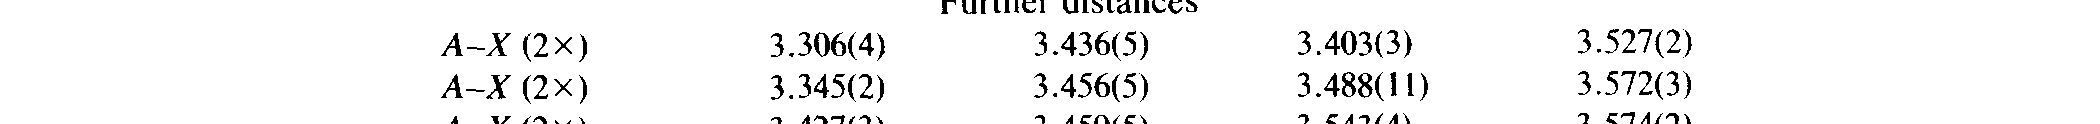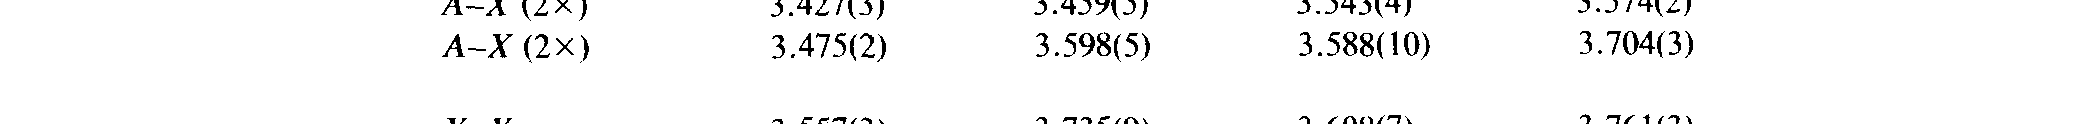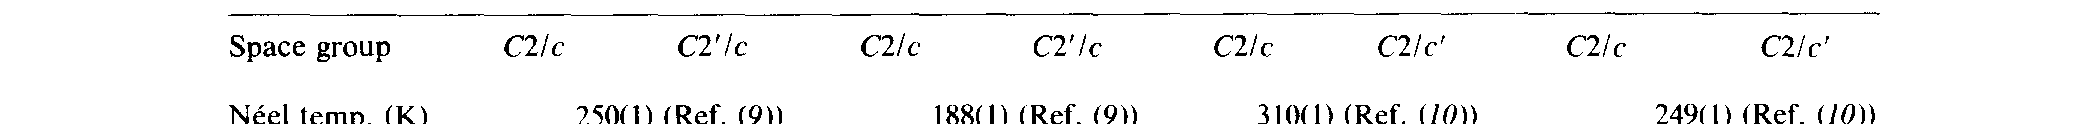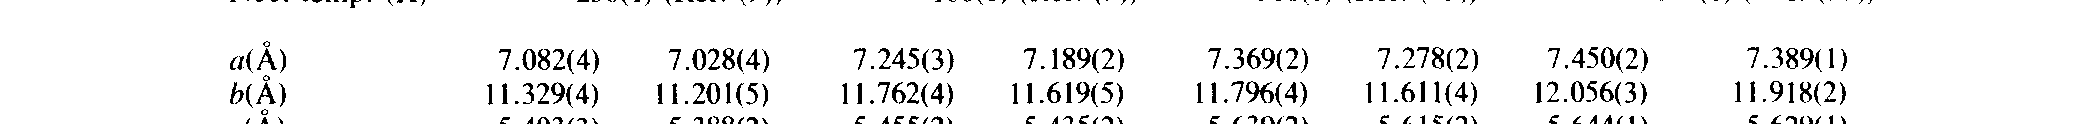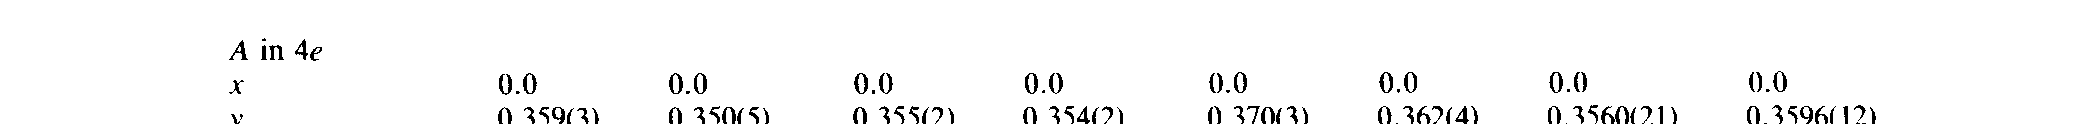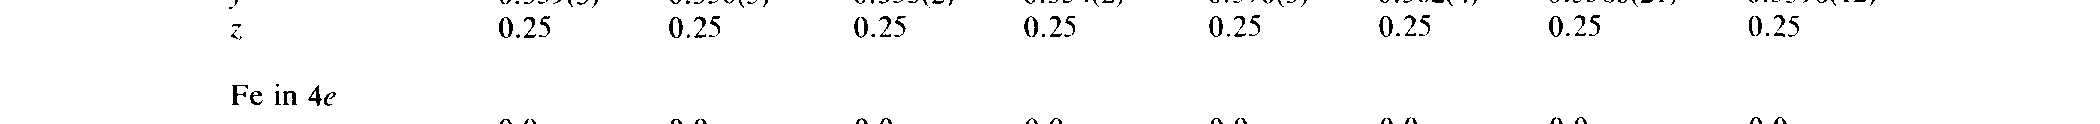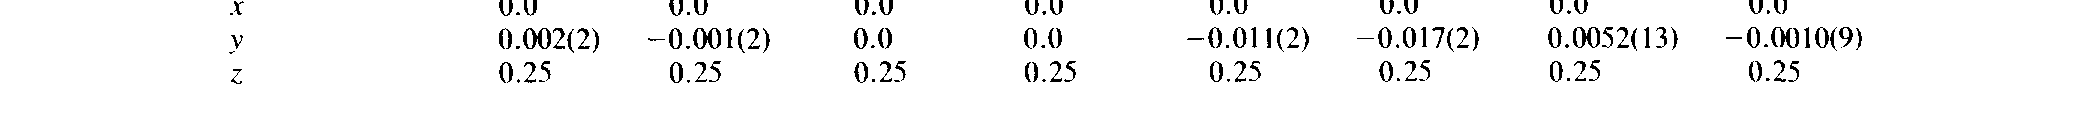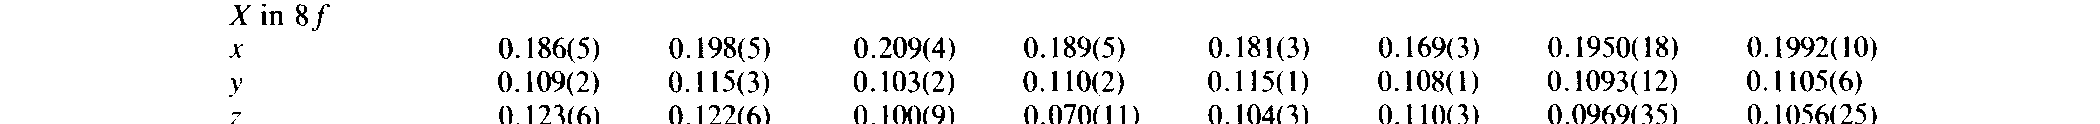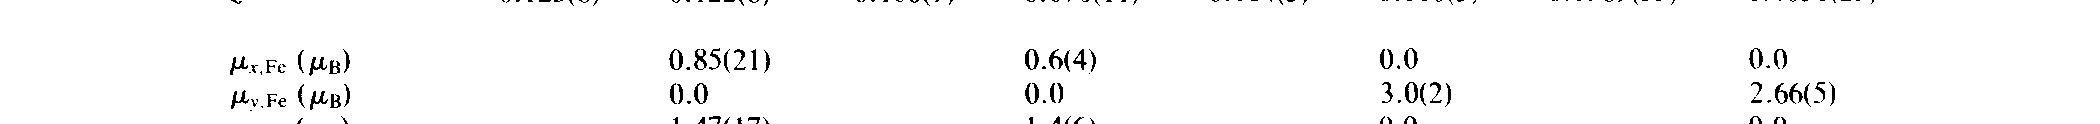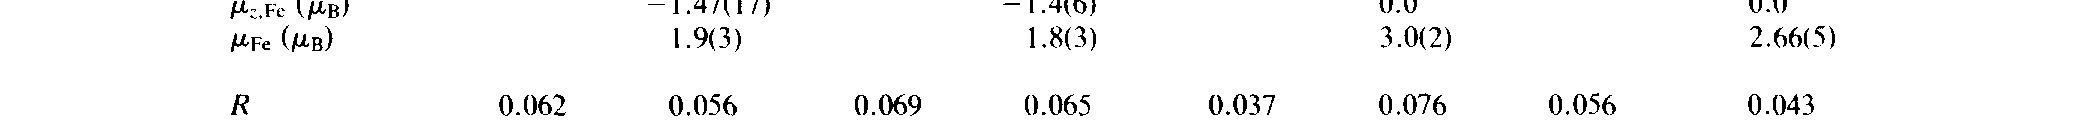⬇ Download Markdown · ⬇ Download LaTeX · ⬇ original · view original · CSV: table 1, table 2, table 3, table 4, table 5, table 6, table 7, table 8, table 9, table 10
JOURNAL OF SOLID STATE CHEMISTRY 70, 262-270 (1987)
The Antiferromagnetic Structures of KFeS2, RbFe$, KFeSez, and RbFeSez and the Correlation between Magnetic Moments and Crystal Field Calculations*
Crystal Field Calculations* W. BRONGER, A. KYAS, AND P. MiiLLER Institut fiir Anorganische Chemie der R WTH Aachen, Prof.-Pirlet-Strasse 1, 5100 Aachen, Federal Republic of Germany Received December 5. 1986
The ternary compounds KFeS?, RbFeS *, KFeSez, and RbFeSe? were prepared and their crystal structures were determined from single-crystal diffractometer data. The atomic arrangement in the isotypic compounds (space group C2/c) is characterized by tetrahedra of chalcogen atoms. Those tetrahedra are centered by iron ions and linked by edges, thus forming chains of L[FeX;2] frameworks (X 4 S or Se). Susceptibility measurements are reported. Neutron diffraction experiments on pbw- dered samples revealed magnetic structures in the antiferromagnetically ordered state. The magnetic moments of the iron ions shows a significant dependence of the atomic arrangement. Therefore calculations based on a simple point charge model are discussed to correlate the magnetic moments and crystal field splittings. o 1987 Academic press, hc.
Introduction KFeSz has been known since 1869 (I, 2). Crystals of KFe& were obtained from the reaction of a mixture of potassium carbon- ate, iron, and sulfur, followed by extracting the solidified melt with water. The structure was first determined in 1942 by Boon and MacGillavry (3). RbFe& and CsFe& were first prepared and their structures determined in 1968 ( 4). Since then the analogous selenium com- pounds have been prepared, too; up to now only powder data have been available for the investigation of their crystal struc- tures (5). The magnetic properties of these alkali chalcogenoferrates are characterized by an- * Dedicated to Dr. H. Nowotny. tiferromagnetic interactions of the iron ions. They have been the subject matter of several publications based on susceptibility measurements and Mossbauer spectra (4, 6-12). Hereafter the results of further ex- periments concerning KFe&, RbFe&, KFeSe*, and RbFeSez are reported. First, detailed single-crystal X-ray data and pow- der neutron diffraction data are given. Fi- nally, the series of magnetic moments is ex- plained by means of the crystal field theory. Thus a further insight in bonding properties is gained. Preparation The compounds KFe& and RbFe!$ were prepared by the reaction of a mixture of alkali carbonate and iron sponge in the mo- lar ratio 4 : 1 in a stream of purified H2S for 0022-4596187 $ 3.00 Copyright 0 1987 by Academic Press. Inc. All righta of reproduction in any form reserved. 262
ANTIFERROMAGNETIC STRUCTURES OF AFeXz 263 4 hr at 1000 K (1.3). The potassium carbon- ate was obtained by dehydration of KzCOl * 1.5 H20 (Alfa-Ventron, ultrapure) in a stream of argon at 420 K. For the rubidium carbonate (Alfa-Ventron) a purity of 99.9% is specified, for the iron sponge (Ziusser) 99.96%. The H2S (Gerling, Holz & Co., pu- rity 99.85%) was dried with calcium chlo- ride and phosphorus(V) oxide. The mixture of alkali carbonate and iron sponge was placed into a corundum boat and then into a quartz tube which was heated up to 1000 K. After the reaction the sample was cooled for 8 hr in an H2S atmo- sphere. Extraction of the solidified melt with water and alcohol yielded well-devel- oped violet needles. Quenching of the sam- ples led to very small crystals, the physical properties of which did not differ from those cooled slowly. Those small crystals were preferred for the experiments with powdered samples. As KFe& and RbFeS are not stable when exposed to air for any considerable length of time, they were stored under argon. The selenides KFeSez and RbFeSe2 were prepared in an analogous manner. The re- action was carried out at 1050 K in a stream of hydrogen and gaseous selenium. For the hydrogen gas (Fa. Linde) as well as for the selenium powder (Johnson Matthey Ltd.), a purity of 99.999% is specified. The reac- tion products were extracted with water and alcohol, dried under vacuum, and stored under argon. Violet needle-shaped crystals similar to the sulfides were formed. Structure Determination The crystal structures of the alkali iron chalcogenides AFeX2 (A & K or Rb, X &! S or Se) were determined by X-ray experi- ments on single crystals. The measure- ments were performed with a computer- controlled single-crystal diffractometer (Nonius CAD4, graphite monochromator). All calculations were carried out using a FIG. 1. Atomic arrangement of KFe&. PDP 11/45 computer with programs of the Nonius SDP system. The results of the final calculations are given in Table I. Table I1 shows the lattice constants, determined by X-ray diffraction on powdered samples (Guinier-Simon method, CuKcv, radiation, a-quartz as standard). The four compounds are isotypic and crystallize in the monoclinic KFeS type (space group C2/c, Z = 4). The atomic ar- rangement is shown in Fig. 1. It is charac- terized by one-dimensionally edge-linked sulfur tetrahedra centered by the iron at- oms. The t[FeS&] chains along the c direc- tion are separated by the alkali atoms. Ta- ble III shows the shortest atomic distances and the X-Fe-X angles in the chains. Susceptibility Measurements Magnetic susceptibilities of the com- pounds KFe&, RbFe&, KFeSez, and RbFeSez were determined by the Faraday method in the temperature range of 3.7-295 K. The samples were sealed in ampoules made from synthetic quartz. The measure- ments at different magnetic field strengths (4 x 10⁵-11 x IO5 Am-‘) showed that susceptibilities depended little on field strength. The Xmol-T diagram (Fig. 2) is based exclusively on values obtained at the strongest field. Diamagnetism was cor- rected by adding the sum of the ionic incre- ments (14). The small positive values of the magnetic
264 BRONGER, KYAS, AND MiiLLER TABLE I X-RAY EXPERIMENTS ON SINGLE CRYSTALS: RADIATION, 20 RANGE, NUMBER OF SYMMETRY-INDEPENDENT REFLECTIONS WITH Fi 2 3 c (Fg) /\ Z( F&-R VALUES, ATOMIC COORDINATES. AND THERMAL PARAMETERS U,, ( 10e4 A2Y - KFeSz RbFeS? K FeSel RbFeSe* Radiation 28 range Z(F;) R value Fe in 4e A in 4e x X in Sf x Y z U,l &2 u33 Ul2 UI3 u23 MoKa Ise535 573 0.03 0.0 0.99668(S) 0.25 176(2) l93(3) 114(2) 0 6W) 0 0.0 0.3572(I) 0.25 283(5) 29 I(6) 270(5) 0 62(4) 0 0.1%0(Z) 0.1098(l) 0.1068(2) 247(3) 239(4) 172(3) -82(3) 9W) -28(3) MoKa lSBS26 337 0.051 0.0 0.9976(3) 0.25 160(10) 230( IO) 79(S) 0 53(7) 0 0.0 0.3553(2) 0.25 237(S) 300(10) 210(7) 0 56(6) 0 0.1905(6) 0.1061(4) 0.0981(7) 250( IO) 250(20) 160( IO) -SO( IO) 70( IO) -2O(lO) AgKff ise524 290 0.075 0.0 -0.0031(6) 0.25 130(20) l50(30) 210(20) 0 70(20) 0 0.0 0.3608( IO) 0.25 380(50) 240(60) 340(50) 0 I40(40) 0 0.2012(5) 0. I l36(3) 0.1099(6) 250( IO) 240( 10) 219(9) - lOO(20) I l3(7) -30(20) AgKa 158524 320 0.03 I 0.0 -0.0021(2) 0.25 217(9) l90(10) 150(S) 0 W6) 0 0.0 0.3585(Z) 0.25 263(7) 291(S) 255(7) 0 I I l(5) 0 0.1948(2) 0.1130(l) 0. IOlS(2) 266(4) 249(5) 194(4) -81(5) I28(3) -38(5) Note. Standard deviations are given in parentheses. u The anisotropic temperature factor is defined as exp[-2+(U,,hza*2 + U~zkzb*z t + 2U,,klb*c*)]. susceptibilities indicate antiferromagnetic can be explained by the change of a one- interactions. There is nothing in these mea- dimensionally to a three-dimensionally or- surements to connect temperatures with dered spin arrangement, consistent with a three-dimensional magnetic ordering, as chain structure. A transition of this kind shown by Mossbauer spectra (9, 10). This does not lead to a significant maximum in
ANTIFERROMAGNETIC STRUCTURES OF AFeXz 265 TABLE II LATTICE CONSTANTS DETERMINED BY THE GUINIER METHOD KFeS* RbFeS? KFeSez RbFeSe? 4‘Q 7.084(3) 7.223(3) 7.342(2) 7.474(2) b(h I I .303(4) I I .725(3) I I .746(2) 12.091(3) c(W) 5.394(2) 5.430(2) 5.629(l) 5.662(2) P(“) 113.2(l) 112.0(l) Il3.52(2) 112.38(2)
the xmol-T diagram, such as is found when Neutron Diffraction Experiments an unordered paramagnetic state changes into an antiferromagnetically ordered one. For these experiments powdered sam- On the other hand, a three-dimensional ples were filled into aluminum tubes of 40 magnetic ordering causes a hyperfine split- mm in length and 8 mm in diameter. The ting in the Miissbauer spectrum and sharp experiments were carried out with the pow- Bragg reflections in neutron diffraction. der diffractometer of the BER II at the TABLE Ill INTERATOMIC DISTANCES (A) AND ANGLES (“) KFeSz RbFe!$ KFeSez RbFeSe, 2.231(2) 2.237(2) Distances and angles in a FeX4 tetrahedron 2.235(5) 2.246(5) 2.363(5) 2.369(6) 2.365(2) 2.367(2) Fe-X (2x) Fe-X (2x) 2.832(l) Fe-Fe (2x) 2.698(l) 2.716(l) 2.815(l) X-X(2X) x-x x-x (2X) x-x 3.561(3) 3.671(4) 3.665(2) 3.756(5) 3.564(9) 3.701(9) 3.683(6) 3.751(S) 3.803(7) 3.865(8) 3.879(5) 3.949(6) 3.790(3) 3.875(3) 3.890(2) 3.939(2) 106.48(4) Il2.8(1) IlO.60(6) X-Fe-X 105.70(5) 114.69(9) ll0.26(8) 105.4(l) I l4.1(3) 110.5(2) Further distances 3.436(5) 3.456(5) 3.459(5) 3.598(5) 107.0(I) I l3.4(3) 110.1(1) A-X(2x) A-X(2x) A-X(2x) A-X(2x) 3.306(4) 3.345(2) 3.427(3) 3.475(2) 3.403(3) 3.488(1 I) 3.543(4) 3.588(10) 3.527(2) 3.572(3) 3.574(2) 3.704(3) x-x 3.557(3) 3.735(9) 3.608(7) 3.761(3) 3.993(6) 4.004(6) 4.274(14) 4.100(2) 4.116(2) 4.360(4) A-Fe (2x) A-Fe (2x) A-Fe 3.876(4) 3.877(2) 4.075(2) 3.978(2) 4.005(4) 4.194(4) 4.315(18) 4.464( 14) 4.441(4) 4.562(3) A-A (2x) A-A (2x) 4.207(2) 4.262(4) 4.346(4) 4.378(5)
266 BRONGER, KYAS, AND MijLLER FIG. 2. Temperature dependence of the magnetic susceptibilities of the compounds AFeX*. Hahn-Meitner Institut in Berlin. Neutron diffraction diagrams were recorded by a po- sition sensitive detector in a 28 range of 80” with a resolution of 0.2” (A = 2.39 A, collimation 10’). Low temperatures were achieved using a closed-cycle refrigerator. High temperatures were kept constant throughout the sample by short-circuiting its far end to the oven using a copper heat conductor. Table IV contains the results of the neu- tron diffraction experiments. The NCel tem- peratures observed in Mossbauer spectra are taken from the literature. Neutron dif- fraction diagrams measured above the Neel temperatures show only those intensities of Bragg reflections which are due to nuclear scattering. Profile-fitting calculations after the Rietveld method were in agreement with the measured profile. Starting values for the refinement were taken from single- crystal X-ray experiments. The neutron dif- fraction diagrams at the lowest tempera- tures showed changes from the intensities of reflections observed at high tempera- tures; and in addition they showed reflec- tions which did not follow the crystallo- graphic extinction h01 only with 1 = 2n. These additional reflections can only be ex- plained by a spin structure with ferromag- netic ordering in the a-b plane and antifer- romagnetic ordering along the c axis. Calculations based on this model led to the parameters given in Table IV after the re- finement through Rietveld’s program (15). For the sulfides a spin structure charac- terized by spins in the a-c plane forming an angle of 20” with the c axis is obtained. In the selenides the spins are aligned parallel to the b axis. This means that the sulfides belong to the Shubnikov group C2’lc, and the selenides to C2/c’. Figure 3 shows the two spin structures. The components of the magnetic moments projected to the crystal- lographic axis and the resulting magnetic moments according to the formula p = 2s~~ are given in Table IV. Previously pub- lished KFeS neutron diffraction data are in agreement with our results (16, 17). Magnetic Moments and Crystal Field Calculations The magnetic moments of the iron atoms in the antiferromagnetic structures of KFeS, RbFe&, KFeSez, and RbFeSez show significant deviations from the value expected for a ds ion with p = 5.0 pa (see Table IV). Obviously spin states with S = 312 a nd S = l/2 play an important part. Our earlier investigations into the mag- netic properties of the ternary sulfide CsFe& revealed low spin states of iron ions. After mechanically destroying linear FIG. 3. Spin structures of AFe& (left side, space group C2’lc) and of AFeSez (right side, space group C2lc’).
ANTIFERROMAGNETIC STRUCTURES OF AFeX2 267 TABLE IV NEUTRON DIFFRACTION EXPERIMENTS: SPACE GROUPS, NOEL TEMPERATURES, PARAMETERS AFTER PROFILE REFINEMENT, AND R VALUES T(K): KFeS? RbFeS? KFeSe? RbFeSe? 29s II 29s II 393 15 295 14 Space group C2lc C2'lc N&A temp. (K) 250(l)(Ref. (9)) 7.082(4) 7.028(4) ll.329(4) 11.201(5) S.403(3) 5.388(2) 113.2(l) 113.3(l) 0.7(S) 0.4(4) A in 4e x 0.0 0.0 I' 0.359(3) 0.350(S) 7 0.25 0.25 Fein4e .r 0.0 0.0 .v 0.002(2) -0.001(2) z 0.25 0.25 Xin8f x 0.186(S) 0.198(S) ? 0.109(2) 0. I U(3) i 0.123(6) 0.122(6) i4FC (ys) 0.85(21) I& Fe (pa) 0.0 kk,FC (PB) -1.47(17) w-e (/%) 1.9(3) R 0.062 0.056 R “UCi 0.05 R m.Agn 0.108 c2ic C2'/c c2/c C2ic' czic CZIC' 188(l) (Ref. (9)) 310(l) (Ref. (10)) 249(l) (Ref. (10)) 7.245(3) 7.189(2) ll.762(4) ll.619(5) 5.455(2) 5.435(2) 112.0(l) 112.2(l) l.l(4) 0.2(4) 7.369(2) 7.278(2) ll.796(4) 11.611(4) 5.639(2) 5.6lS(2) 113.5(l) 13.7(l) 5.0(3) 0.4(4) 7.450(2) 7.389(l) 12.056(3) 11.91X(2) 5.644(l) 5.629(l) 112.39(l) 112.55(l) 0.9(4) 0.6(3l 0.0 0.0 0.0 0.0 0.0 0.0 0.355(2) 0.354(2) 0.370(3) 0.362(4) 0.3560(21) 0.3596(12) 0.25 0.25 0.25 0.25 0.25 0.25 0.0 0.0 0.0 0.0 0.25 0.25 0.0 0.0 -0.011(2) -0.017(2) 0.25 0.25 0.209(4) 0.1890) 0.103(2) 0. I lO(2) O.lOO(9) 0.070(11) 0.6(4) 0.0 -1.4(6) 1.X(3) 0.069 0.065 0.037 0.076 0.061 0.068 0.127 0.124 O.l81(3) 0.169(3) 0.115(l) 0.108(l) 0.104(3) O.llO(3) 0.0 3.0(2) 0.0 3.0(2) 0.0 0.0052(13) 0.25 0.1950(18) 0.1992(10) 0.1093(12) 0.1105(6) 0.0969(35) 0.1056(25) 0.056 0.0 -0.0010(9) 0.25 0.0 2.66(S) 0.0 2.66(T) 0.043 0.038 0.101
magnetic interactions in the t[FeS4i2] chains of &Fe&, the iron ion was found to be in a low spin state. Furthermore in mixed crystals of CsGar-, Fe& measure- ments of magnetic susceptibilities pointed to a low spin configuration of Fe3+ (6). This conforms to the low magnetic hyperfine splitting in the Mossbauer spectrum point- ing to nearly complete spin pairing (9). These anomalous magnetic moments of transition metal ions observed here in the tetrahedral holes of a framework structure can be explained by means of the crystal field theory. This theory becomes applic- able mainly for the reason that in the tern- ary alkali transition metal chalcogenides AxMJz the strong electropositive alkali metal atoms build up a pressure of elec- trons, thus forming complex [M,Xz]“- an- ions. Therefore a three-dimensional delo- calization of electronic states loses some of its importance. When x is increased, thereby raising the electronic pressure in the lattice, disintegration of the [M,XJ- frameworks in the ternary compounds A,M,X, is to be expected (18). The chalco- genides AFeX2 discussed here contain one- dimensional [Fe&- and [FeSe$ chains of edge-linked tetrahedra, Na3FeS3 (I 9) [Fe2S,16- double tetrahedra, and finally
268 BRONGER, KYAS, AND MiiLLER Na5FeS4 (20), which is richest in alkali metal isolated [FeS415- t etrahedra. We will include the isolated [Fe&]“- groups to- gether with the chain structures when dis- cussing the results of our crystal field calcu- lations to achieve an extended model. As a first step in a perturbation calculation the oxidation numbers of the atoms concerned were taken as point charges to assess the influence of the coordinating atoms on the electronic states of the central iron atom. Details of this procedure, which follows the strong field method, have been described in various papers. (18, 21, 22). In Fig. 4a the energy splittings of the d orbitals of a Fe3+ ion are shown, first, in a regular tetrahedral coordination of S*- an- ions, and second, in real coordination, which is found in Na5FeS4 and includes only the four nearest sulfur atoms (r < 2.3 A). Further splittings are pointed out, in- cluding the influence of all ions up to the specified radius r around the central atom. The result shows that ions farther away than the first ligand sphere essentially do not change the splitting of the d orbitals. The [FeSJ- tetrahedra can be regarded as swimming in a sea of sodium ions. This sit- uation is quite similar to that of a solution of complex ions. El When computing analogous diagrams for the structure of Na3FeS3 the picture changes drastically (cf. Fig. 4b). In this case the Fe3+ ion occupying the neighbor- ing tetrahedral hole in the [Fe2S$ unit at a distance ~3.0 A exercises a powerful influ- ence on the energy splitting of the d orbit- als. This influence is diminished with regard to those ions which are farther away from the central atom (up to 5.4, 8.0 A); the re- maining energy splitting; however, is dis- tinctly larger than that arrived at for the isolated tetrahedron (Fig. 4a). The influ- ence of a higher number of atoms than those belonging to the first ligand sphere is even more prominent in the chain struc- tures of the compounds KFeS2, RbFeSz, KFeSe2, and RbFeSe? (cf. Fig. 4~). Those d states participating in the z direction are ac- cordingly reduced in energy, as the positive charges of the iron atoms are arranged at a relative short distance in the same direc- tion, i.e., along the chain. In Fig. 5 the results of the crystal field calculations are compared. In the series of splitting diagrams there is an unmistakable correlation of the measured magnetic mo- ments and the energies computed on a rela- tive scale. Small energy splittings as calcu- lated for Na5FeS4 c orrespond to a magnetic E - -- --- - -- --- = -I --=- --= - --_--- -- - - - - --- ---_-__- = _- ---- --_- -- -_---- rid, 23 3.0 ‘.3 5.‘ 8.0 ao 12.0 u.0 23 27 ‘3 5.L 8.0 x1.0 v.0 1‘0 bl cl FIG. 4. Energy splittings in dependence of point-charge model calculations including different num- bers of ions. (a) NaSFeS.,, (b) Na3FeS,, (c) KFe&.
ANTlFERROMAGNETlC STRUCTURES OF AFeXz 269 E t - - - - - - - - I - I - I =3 - = = - i - = = - I - - - - 9 e CSFKl‘ NoskS NqFeS3 we52 -52 wese2 Rbwq pbgl 50 ‘7 - ,911, 1.8131 3.0 121 26615, FIG. 5. Energy splittings in comparison with the magnetic moments. moment for the Fe’+ ion approaching a high spin value. In the case of the largest energy splittings, calculated for KFe& and RbFe&, the moments were found to be spread between a low spin value (S = l/2) and a medium spin value (S = 3/2). The chain structures of the selenides follow the same pattern. The computed energy split- tings are smaller than those for the sulfides, but distinctly larger than for an isolated tet- rahedron. Thus magnetic moments larger than those in the sulfides but smaller than a high spin value are to be expected. Neutron diffraction experiments confirm this. For the compound Na3FeS3 we have as yet not been able to determine the magnetic mo- ments from neutron diffraction experi- ments; nevertheless a magnetic moment distinctly smaller than 5.0 pa is to be ex- pected. The use of oxidation numbers as point charges in our calculations naturally leads to a maximum value of energy splittings. Analogous calculations with reduced point charges showed smaller splittings as was to be expected. Nevertheless the relative se- quence discussed is valid. The increase of Fe-S distances in the iso- lated [FeS415- g roup with regard to those in the [Fe&- chains corresponds to the series of magnetic moments: For iron ions in the low spin state a smaller radius than for those in the high spin state is to be ex- pected. The deviations of the measured magnetic moments from those calculated for the dif- ferent spin states can be explained by the neglect of interelectronic interactions and spin-orbit coupling. Interactions of excited states with ground states may be respon- sible for the deviations from spin-only val- ues. In Fig. 5 the halogenide CsFeCL, (23) is alongside the chalcogenides. Susceptibility measurements have revealed a magnetic moment equaling the spin-only value for a Fe3+ ion with five unpaired electrons. This compound contains isolated [FeCl& an- ions; their spectroscopic data as measured in solution are available. In the literature a AE value for the splitting of the e and t7 state of about 5000 cm-’ is quoted (14). Transferring this value into the series of splittings in Fig. 5, spin-pairing can be ex- pected for those compounds with linked tetrahedra. Spectroscopic investigations
270 BRONGER, KYAS, AND MULLER will be carried out in due course. However, the above-mentioned relative calculations already convey a detailed picture which also has been observed in a series of ternary manganese compounds (21, 24). Some months ago extended Htickel cal- culations for the [Fe&- chains of our com- pounds were published by Silvestre and Hoffmann (25). With regard to the splitting of d orbitals their analysis is not inconsis- tent with our results. The advantage of the model as described above is that it pro- duces information on local properties in a simple approximation, thus allowing pre- dictions with regard to other compounds. Acknowledgments The authors express their thanks to Mr. K. Kruse for performing the single-crystal diffraction experi- ments. Thanks are also due to Prof. Dr. H. Dachs for giving access to the multicounter at the BER II in the Hahn-Meitner Institut in Berlin. Moreover, we are grateful to the Bundesminister fur Forschung und Technologie and the Fonds der Chemischen Industrie for financial support. References I. K. PREIS, J. Prukt. Chem. 107, 12 (1869). 2. R. SCHNEIDER, J. Prakt. Chem. 108, 16 (1869). 3. J. W. BOON, AND C. H. MACGILLAVRY, Rec. Z’ruu. Chim. Pays-Bas 61, 910 (1942). 4. W. BRONGER, Z. Anorg. Allg. Chem. 359, 225 (1968). 5. W. BRONGER, Naturwissenschuften 53, 525 (1966). 6. W. BRONGER, AND P. MUELLER, .I. Less-Common Met. 70, 253 (1980). 7. P. MOLLER, dissertation, Technische Hochschule Aachen (1980). 8. A. KYAS, dissertation, Technische Hochschule Aachen (1981). 9. H. P. NISSEN, AND K. NAGORNY, Z. Phys. Chem. NF 95, 301 (1975). IO. H. P. NISSEN, AND K. NAGORNY, Z. Phys. Chem. NF 99, 14 (1976). II. W. V. SWEENEY, AND R. E. COFFMAN, Biochim. Biophys. Acta 286, 26 (1972). 22. D. C. JOHNSTON, S. C. MRAW, AND A. J. JACOB- SON, Solid State Comm. 44, 255 (1982). 13. D. SCHMITZ, AND W. BRONGER, Nuturwi.ssen- schaften 58, 322 (1971). 14. A. WEISS, AND H. WITTE, “Magnetochemie,” Verlag-Chemie, Weinheim (1973). 15. H. M. RIETVELD, J. Appl. Crystallogr. 2, 65 (1969). 16. T. TOMKOWICZ, A. SZYTULA, AND H. BAK- P~ASIEWICZ, Phys. Status. Solidi A 57, K25 (1980). 17. M. NISHI, AND Y. ITO, Solid State Commun. 30, 571 (1979). 18. W. BRONGER, AND P. MUELLER, J. Less-Common Met. 100, 241 (1984). 19. P. MUELLER, AND W. BRONGER, Z. Naturfbrsch. B 34, 1264 (1979). 20. K. 0. KLEPP, AND W. BRONGER, Z. Anorg. Ally. Chem. 532, 23 (1986). 21. W. BRONGER, Pure Appl. Chem. 57, 1363 (1985). 22. A. L. COMPANION, AND M. A. KOMARYNSKY, Chem. Educ. 41, 257 (1964). 23. G. MEYER, Z. Anorg. Allg. Chem. 436, 87 (1977). 24. W. BRONGER, R. H~PPNER, P. MUELLER, AND H.-U. SCHUSTER, Z. Anorg. A&. Chem. 539, 175 (1986). 25. J. SILVESTRE, AND R. HOFFMANN, Inorg. Chem. 24,4108 (1985).